GMP Audit Readiness Guide for Pharma and Life Sciences Teams

In pharmaceutical manufacturing, an audit can be a routine event on the calendar or a surprise knock at the door. Either way, whoever owns your GMP compliance has to be ready. And whether you are preparing for a WHO, FDA, or MHRA audit, readiness is how you protect patients, safeguard product quality, and prove to regulators that your operations are compliant from the inside out.
This guide is written for the people doing the work: quality leads, lab managers, operations and maintenance, production, and leadership — the ones who do not have time to decode acronyms or read a 200-page PDF to get audit-ready. It maps what readiness looks like in practice, on the floor, who is involved, where things get missed, and how to avoid the problems that usually get flagged when regulators arrive.
What are GMP audits?
A GMP (Good Manufacturing Practice) audit is a structured review of your facility, processes, documentation, and people, carried out to confirm your operations meet quality and safety standards. These are not paperwork checks. They are deep dives into whether you can consistently produce medicines that are safe, effective, and fully traceable. In short, the regulator is asking a single question: can we trust what you are doing to meet the required standards and operate to best practice?
Audits can be triggered in several ways:
- Routine regulatory cycles, such as MHRA, FDA, or WHO inspections
- Product quality complaints or adverse event reports
- New facility validations or post-approval changes
- Internal QA and compliance assessments
Whether it is a global authority or an internal QA manager walking the floor, readiness shows you have built a culture of accountability rather than panic.
Who needs to be GMP-ready?
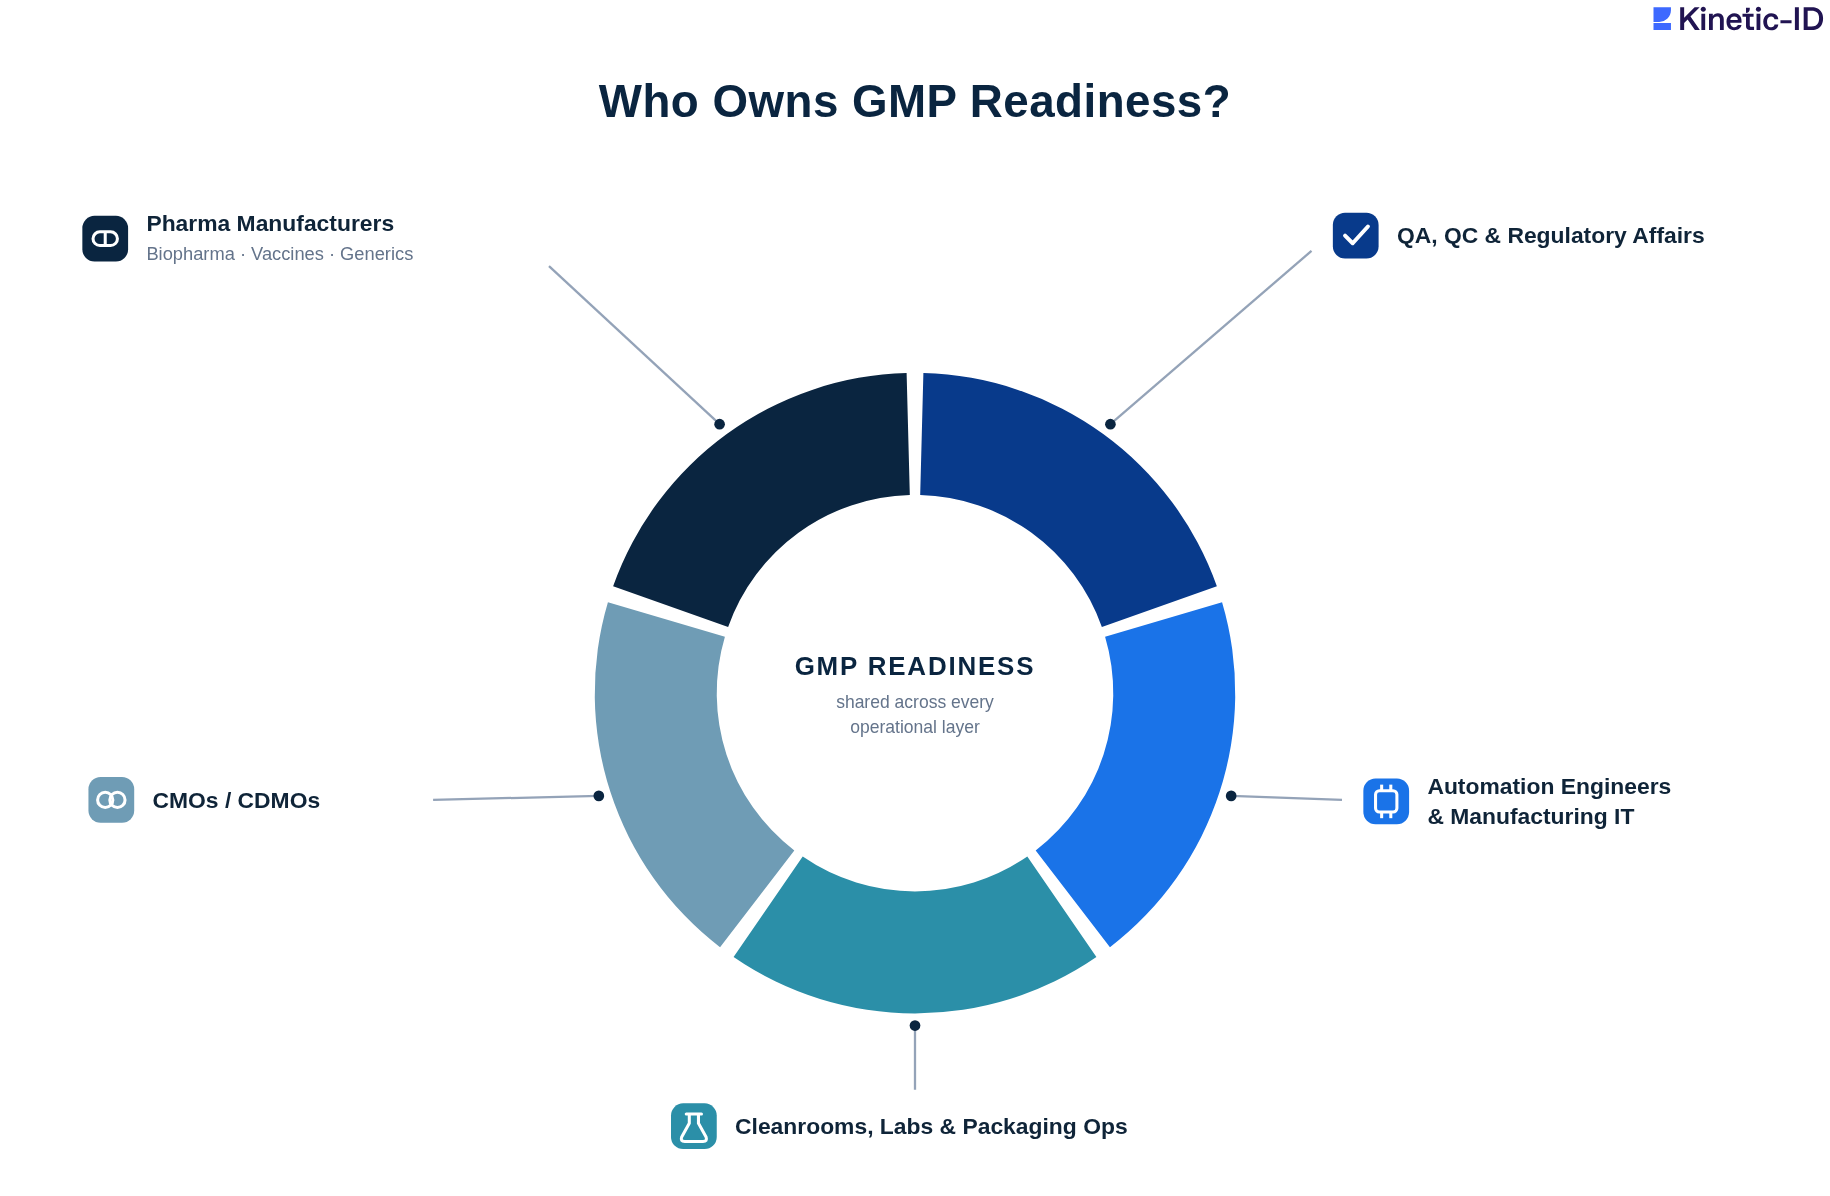
GMP compliance involves far more than QA. It is a team effort, and the teams that should actively prepare include pharmaceutical manufacturers across biopharma, vaccines, and generics; Contract Manufacturing Organisations (CMOs); cleanroom, lab, and packaging operations; automation engineers and manufacturing IT teams; and QA, QC, and regulatory affairs personnel. Whether you run a GMP facility, a Grade A cleanroom, or a Laboratory Information Management System (LIMS), everyone has a role in maintaining a compliant environment. The most resilient sites treat readiness as a shared responsibility, with ownership assigned at every operational layer.
What does GMP readiness cover?
This is where many teams lose track. GMP is not just about documents or tidy floors. A compliance and adherence checklist should cover six areas, each of which feeds directly into your audit trail.
Use these categories to design an internal checklist that mirrors the structure of common audit frameworks, so your own review runs along the same lines an inspector will.
Where does GMP readiness matter most?
Not all areas attract equal scrutiny. Concentrate on GMP and GxP facilities, classified cleanrooms (Grades A, B, C, and D), controlled but non-classified areas such as labs, packaging zones, and warehousing, and MES/LIMS-controlled production rooms. Critical areas like gowning rooms, sample storage, and packaging zones need to be monitored as diligently as your primary manufacturing lines, because a weak link in any of them can undermine the whole facility's compliance. Much of this comes back to visibility — the ability to see what is happening where the work is — a theme explored in If You Can't See It, You Can't Fix It.
When should you prepare?
Waiting for an audit letter is too late. Readiness should be built into operations year-round, and especially before scaling production or launching new products, ahead of site expansions, renovations, or equipment installs, in advance of scheduled or surprise audits, and before digitisation efforts such as MES, LIMS, or eBatch Record rollouts. Treating readiness as a continuous process moves a team from reactive mode into a culture of proactivity, and monthly walkthroughs paired with annual mock inspections de-risk the unknowns. The same continuous mindset underpins Driving Continuous Optimization in Life Sciences Manufacturing.
Why does it matter?
GMP is not about ticking boxes to get through an inspection. It is about embedding quality and control into everyday processes, and when teams treat compliance as an afterthought, the consequences are serious and immediate:
- Audit findings and compliance warnings, such as FDA 483s and WHO observations
- Product recalls and stock quarantines
- Regulatory fines, licence suspension, and reputational fallout
Readiness, in other words, is risk mitigation — an operational insurance policy that protects patients, products, and the business.
How Kinetic-ID actually helps with GMP audit readiness
Staying GMP-compliant is not about the tidiest documentation binder. It is about what actually happens day to day, shift to shift, on the floor rather than in a filing cabinet — and that is where many teams hit a wall. The processes exist, but the tools often are not built to keep up.
Kinetic-ID's mobile workstations were built because operators kept describing the same problem: they could not reach facility data, real-time information, floor visibility, or SOPs without breaking gowning or operating procedures, and they lost time walking between cleanroom zones just to log a deviation. Bringing the systems to the point of work closes that gap:
- Facility visibility and access to multiple data sources, matched to your IT/OT infrastructure, with full mobility across sites.
- Compliance without breaking rhythm. Stainless steel and smooth, easy-to-clean surfaces mean hygiene is designed in rather than worked around.
- Everything at the cart. SOPs, LIMS, quality and document management, and MES are all accessible where the work happens, so operators do not have to leave the room or guess the next step.
- Real-time logging. Scanners, label printers, and data logging are built in, which reduces gaps and removes the "we'll fill it out later" slips that become findings.
- Ergonomics that hold up. Comfortable and adjustable, because a 12-hour shift should not force a choice between compliance and the operator.
Each workstation is specified for a particular class of environment, so the hardware is part of the validated chain rather than tech bolted onto a trolley.
The wider case for treating hardware as part of readiness is set out in How to Choose a GMP-Compliant Cart, and the link between point-of-work access and fewer findings in How to Reduce Deviations in Life Sciences Manufacturing.
The 5 Ps of GMP
A useful shorthand for what an audit examines is the five Ps. Weaken one, and the others start to wobble.
Frequently asked questions
Work out what continuous audit readiness is worth on your line
The cost of a findings-heavy audit is rarely one number. Put your own figures against it — audit cycle, deviation and rework cost, operator hours lost to logging away from the point of work, and batch value at risk.
Estimate your ROI